



A molecular simulation box can look like a setup detail—until periodic boundaries make your solute interact with one of its own copies. That can distort the forces and undermine the simulation. The practical challenge is to choose a box that keeps periodic images far enough apart without filling unnecessary space with solvent.
Here is a useful way to think through that choice when preparing a system in GROMACS Wizard.
First, know what periodic boundaries imply
With periodic boundary conditions, the simulation cell is repeated in every direction. For short-range non-bonded interactions, GROMACS uses the minimum-image convention: it considers the nearest periodic image of each particle. Long-range electrostatics need additional treatment, such as PME or Ewald methods, but the box still has to meet the relevant cutoff restrictions.
For a quick setup check, use the documentation’s practical rule of thumb: leave at least 1.0 nm between the solute and the box boundary. That gives at least 2.0 nm between periodic images of the solute. Treat this as a starting point, not a replacement for checking the cutoff restrictions for your simulation.
Choose a shape that suits the solute
A cubic box is familiar, but it is not always the most economical choice. For an approximately spherical macromolecule in solvent, a rhombic dodecahedron or truncated octahedron can fit the solute more efficiently. These shapes are closer to spherical than a cube, so they can use fewer solvent molecules while maintaining the same image distance.
The rhombic dodecahedron is the smallest and most regular space-filling unit cell described here. At the same image distance, its volume is about 71% of a cube’s volume. For spherical or flexible molecules in solvent, the documentation notes that this can save about 29% of CPU time.

Other supported choices include cubic, orthorhombic, and triclinic cells. The best shape depends on the system and setup; a more compact shape is not automatically the right choice for every solute.
Two ways to size the box
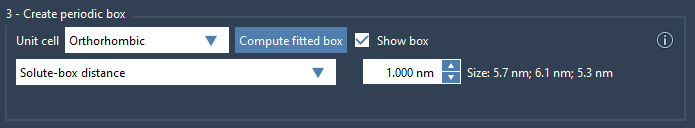
- Box lengths: Set the box dimensions directly. The initial fit is tight, so increase the dimensions as needed to meet the minimum-image requirement. This option also lets you position the system within the box. For a batch project, it can keep the box size consistent across conformations or a path.
- Solute-box distance: Set the distance between the solute and the box. The documentation recommends at least
1 nmas a practical way to help satisfy the minimum-image convention. In a batch project, each conformation or path frame can have a different box size.
A quick setup check
- Consider the overall shape of your solute. For an approximately spherical molecule, compare the rhombic dodecahedron and truncated octahedron with a cube.
- Choose whether you need fixed box dimensions or a set distance from the solute to the box.
- Check the distance to the box boundary and confirm that the resulting periodic-image separation and interaction cutoffs are appropriate for your simulation.
- If you load GROMACS results later, remember that GROMACS stores particles in a brick-shaped volume for efficiency. SAMSON attempts to detect the unit-cell type when importing trajectories, and the type can be adjusted in the importer dialog.
The key is not simply to make the box large. It is to give the solute enough separation from its periodic copies while choosing a cell shape and sizing method that fit the system you are studying.
For the full explanation and links to related setup steps, see the GROMACS Wizard guide to periodic boundary conditions.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.