



A molecular system can look ready for simulation and still be poorly prepared if its periodic box is too small. Under periodic boundary conditions, the simulation treats the system as if copies of it repeat in every direction. If a protein is too close to its periodic image, those copies can interact in ways you did not intend. The challenge is to give the solute enough room without guessing at box dimensions or overlooking how your choice affects a batch of structures.
In SAMSON’s GROMACS Wizard, the box setup is part of preparing the system for simulation. Here’s a practical way to think through the choices before adding solvent or ions.
First, decide what needs to fit
Select the structures you intend to prepare in the Document view, or leave the selection empty if you want to prepare the whole system in the document. The box must include every structure selected for preparation. This matters especially when a document contains multiple models: fitting a box around only one of them may not be appropriate if you later prepare several.
GROMACS Wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated octahedron unit cells. For a straightforward starting point, the tutorial demonstrates an orthorhombic box. Choose the unit-cell type, click Compute fitted box, and select how the box should be fitted.
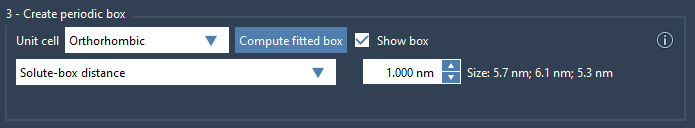
Choose the fitting method based on your project
Solute-box distance sets a distance between the system and the box boundary. The documentation recommends at least 1 nm to help satisfy the minimum image convention. This is a useful option for a single structure when you want to specify the amount of space around the solute.
Box lengths lets you specify the box dimensions directly. The fitted box initially fits tightly around the system, so increase its size as needed to provide adequate separation from periodic images. This option can also be useful for batch preparation: the initial box is determined from all conformations or the full path, helping keep one box size for the batch.
For a batch project, the choice has a practical consequence. With Box lengths, the initial size is based on the whole set of conformations or path. With Solute-box distance, each conformation or trajectory frame can have a different box size. Choose according to whether consistent box dimensions or per-structure fitting better suits the batch you are preparing.

Use the minimum image convention as a check
The key question is not simply whether the molecule fits inside the box; it is whether it has enough space from its periodic copies. The tutorial recommends at least 1.0 nm between the system and the box edge. In its example, that corresponds to at least 2.0 nm between periodic images of a protein or other non-solvent molecules. A molecule may change conformation during a simulation, so consider the system’s possible extent—not only how compact it looks in the starting structure. For a custom force field, check its publication for any relevant cutoff guidance.
After computing the fitted box, you can adjust its unit-cell parameters. Before moving on, confirm that all selected structures are included and that the box has the intended clearance. A fitted box is a starting point, not an automatic guarantee that your chosen size is suitable for every simulation.
Only then add solvent and ions
If you want explicit solvent, choose a water model supported by the selected force field and enable Add solvent. Choosing “none” prevents adding solvent and ions. Ions replace water molecules, so solvent is required if you plan to neutralize the system or add salt. GROMACS can automatically add ions to neutralize the system; additional ions can also be specified by number or salt concentration.
These choices belong together: box dimensions affect the cell, and the cell volume is used when calculating how many ions to add for a specified salt concentration. Check the selected solvent and ion options as part of the same preparation review, rather than treating them as unrelated finishing touches.
Before clicking Prepare: confirm the selected structures, unit-cell type, fitting method, clearance from periodic images, water model, and ion settings. The prepared system can then be imported into SAMSON for inspection before continuing to energy minimization.
For the complete preparation workflow, including force fields, custom topology files, and batch projects, learn more in the GROMACS Wizard preparation documentation.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.