



Two proteins can look broadly similar and still differ in the part you need to compare. A whole-structure superposition may make the overall relationship easier to see, but it can leave a particular helix, binding-site region, or other selected segment poorly aligned. That can make it difficult to answer a practical question: are these specific regions structurally alike?
A useful approach is to treat global and region-specific alignment as two different views of the same comparison. First align the proteins as a whole. Then select corresponding residues and align again using only that selection. The second result focuses the superposition on the region of interest rather than asking the entire structures to fit at once.
Start with a whole-protein comparison
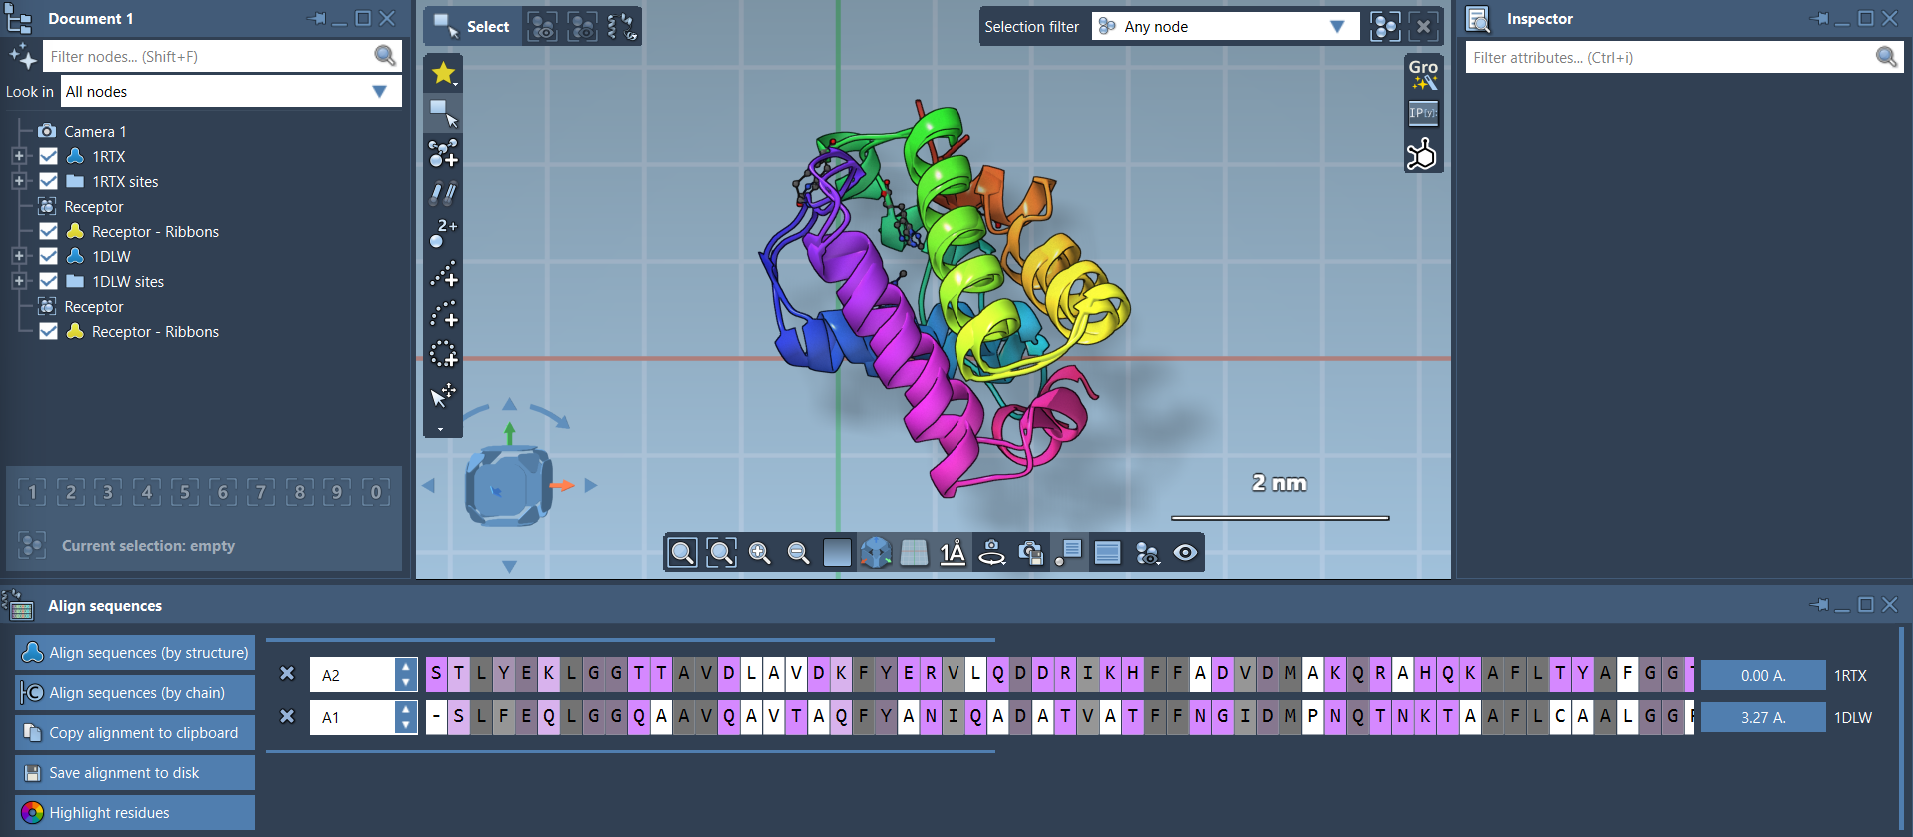
In SAMSON, fetch or open the structures you want to compare, then launch Home > Align to open Protein Aligner. The documentation example uses hemoglobins 1DLW and 1RTX; the same workflow can be applied to other proteins. If your structures contain solvent or ligands you do not want included in the comparison, prepare them first using the protein preparation workflow.
Protein Aligner can align sequences by structural model or by chain. The model option compares models, while the chain option is useful when a model contains multiple chains, such as in an oligomer. Sequence alignment gives you a way to inspect corresponding residues before you superimpose the structures.

As you inspect the alignment, hovering over a residue shows its name and ID and highlights it in the 3D viewport. Clicking a residue selects it and its aligned partners. This connection between the sequence view and the structure view helps you check that the residues you intend to compare correspond across the proteins.
Use the global fit as a reference
To superimpose the full structures, make sure no residues are selected, then click Align to this on the row of the model you want to use as the reference. The other model’s button reports the RMSD for the resulting fit. Treat that value as a summary of this particular alignment, not as a complete description of every local region: a single global fit can distribute differences across the full structure.
Refocus the fit on a region
If a segment looks similar but is not well superimposed, return to the sequence alignment and select the corresponding residues in both proteins. In the tutorial example, the first 20 residues are selected in both sequences to focus on two alpha-helices. Once the selection is made, click the alignment button next to that selection. The structures are then superimposed using those selected residues.

This local fit is useful when your question is about a specific region rather than the overall fold. Compare the selected-region result with the whole-protein result: the contrast can show whether the local segment has a different structural relationship from the proteins as a whole. Be clear about which residues were used, since changing the selection changes what the superposition represents.
A practical checklist
- Choose whether the comparison should be by model or by chain.
- Inspect the sequence alignment and verify the corresponding residues.
- Run a whole-protein superposition with no residues selected to establish a reference.
- Select the region of interest in both sequences, then align using that selection.
- Interpret each RMSD in the context of the residues used for that fit.
This two-pass workflow avoids relying on a global fit when the scientific question is local. It also makes the comparison easier to communicate: you can show both the overall superposition and the focused alignment, with the selected region made explicit.
For the complete step-by-step tutorial, see Align protein sequences and structures in SAMSON.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.