



A molecular system can look perfectly prepared in a viewer and still be poorly set up for simulation. One common source of trouble is the periodic box: make it too small and the system may interact with its own periodic images; make choices inconsistently across a batch and the resulting systems may not be comparable in the way you intended.
The box is not just empty space around a molecule. Under periodic boundary conditions, copies of the simulation cell tile space. The goal is to give the solute enough room that it does not encounter its periodic image in a way that violates the minimum image convention.
Choose the box workflow that fits your project
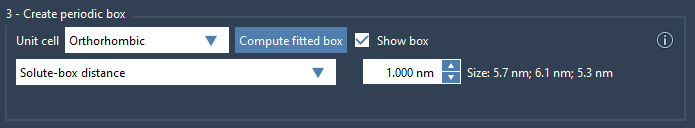
In GROMACS Wizard, you can fit a box using either Solute-box distance or Box lengths. The right choice depends on whether consistent dimensions or a consistent clearance around the solute matters more.
- Solute-box distance: specify the distance between the system and the box. The documentation recommends at least 1 nm. This is a useful starting point when you want a defined margin around the solute. For batch projects, the box size can differ for each conformation or trajectory frame.
- Box lengths: specify the dimensions of the box. Fitting initially makes the box tight around the system, so you need to increase its size as needed. For batch projects, the initial dimensions are determined from all conformations or the full path. This option is useful when you want the same box size throughout the batch.
These options answer different questions: How much clearance should surround each structure? versus What dimensions should every system use? Decide based on the goal of the simulation, rather than choosing solely for convenience.
A practical setup sequence
- Choose what the box must contain. Select the molecular system in the Document view, or leave the selection empty to prepare the whole system in the document. For multiple structures, make sure the selected structures are the ones you intend to prepare.
- Select a unit-cell shape. GROMACS Wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated octahedron cells. The tutorial demonstrates an orthorhombic cell; the appropriate shape depends on your system and simulation setup.
- Fit the box, then inspect its dimensions. Click Compute fitted box and choose the fitting option that matches your project. Fitting determines a starting position and size from the system; it does not remove the need to check that the box provides adequate space.
- Check the clearance. With box lengths, increase the dimensions to leave enough space between the solute and box edges. The documentation recommends at least 1.0 nm of distance. This corresponds to at least 2.0 nm between periodic images of a protein or other non-solvent molecules. Conformations can change, so consider the system’s possible range of motion, not only its starting structure.
- Confirm every selected structure fits. The box must include all structures selected for preparation. This matters especially for a batch project: one conformation may extend farther than another.

Why the margin matters
With periodic boundaries, a protein can effectively encounter copies of itself across the cell. If the box is too small, those encounters can affect calculated forces. The minimum-image requirement is therefore a physical setup consideration, not merely a visual preference. The 1 nm recommendation is a practical guide in the tutorial, but if you use a custom force field, check the relevant force-field guidance as well.
If you plan to add explicit solvent or ions, choose a water model and enable solvent addition during preparation. Ions replace water molecules, so solvent must be present if you want to add ions. Box setup, solvent, and ion choices are connected parts of preparing the system—not independent finishing touches.
After preparation, review any warnings and the output log. If you load the prepared system into SAMSON, you can inspect it in the viewport and document view. That final visual check can help catch an obvious mismatch between the intended system and the structure that was prepared, though it does not replace checking the box dimensions and simulation parameters.
For more detail on the preparation workflow and periodic-boundary considerations, see the GROMACS Wizard preparation tutorial.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.