



A protein–ligand simulation can stall before the first dynamics run—not because the binding pose is wrong, but because the ligand’s structure and topology do not agree. Atom names, hydrogens, residue names, and force-field files all need to line up. A little organization before opening the preparation workflow can prevent a confusing round of troubleshooting.
This guide focuses on the handoff: turning a ligand into a parametrized component, bringing it together with the protein, and giving GROMACS Wizard the files it needs. It covers non-covalent complexes. Covalently linked ligands need additional treatment for the bond connecting the ligand and protein.
Start with a consistent ligand structure
If the ligand is not already a standard residue in the force field you plan to use, it needs its own parameters. Choose a parametrization tool that supports that force field: for example, Antechamber for AMBER, ATB for GROMOS96 54A7, CGenFF for CHARMM, or LigParGen for OPLS-AA. The key is to choose the force-field approach first, rather than generating a topology and only later discovering it does not fit the intended setup.
Many parametrization services require the ligand to have hydrogens. In SAMSON, use Edit > Add hydrogens. For ligands in the Chemical Component Dictionary (CCD), SAMSON can add hydrogens with the CCD names. For other molecules, hydrogen placement is based on valences, so check that aromaticity and charges are already represented correctly. If the ligand file contains useful chemical information, such as a .mol2 file, use that as the input when adding hydrogens.
If the ligand is inside a complex, select it in the Document view and use Home > File > Save selection as… to export it in a format accepted by the chosen service. Before exporting, check the service’s accepted formats. If you already have a suitable hydrogenated ligand file, you can use it directly.
Keep the structure and topology in agreement
After parametrization, the essential deliverable is usually an include topology file (.itp). The service may also provide an updated structure file and, in some cases, a custom force field. Keep these outputs together: the updated structure is useful for checking whether atom names or hydrogens changed, while the topology and any required force-field files are needed for preparation.
One small detail has outsized importance: the ligand name in the structure must match the name used in the topology. If you combine files from different sources or replace a ligand in an existing complex, verify that the names still agree. Also compare the updated ligand structure with the one in the complex. If parametrization changed atom names or added hydrogens, the original ligand may no longer be an exact match.
If you need to replace the ligand already present in the complex, position the updated ligand at the old ligand’s location, then delete the old one. If you are assembling a complex from separate files, you can place the ligand with SAMSON’s move tools, dock it with AutoDock Vina Extended, or append it to the protein structure file. When appending, make sure the structures do not overlap and are arranged in separate chains or with consecutive sequence numbering.
Give GROMACS Wizard the required files
Open GROMACS Wizard via Home > Apps > Biology > GROMACS Wizard, switch to Prepare, select the complex in Document view, and choose Set system.

In the Choose model section, use Edit for include topology files (itp) and browse to the ligand’s .itp file. If that topology relies on a custom force field not available by default, download it from the parametrization service and add it using the force-field Add button. For example, ATB may provide a force field such as gromos54a7_atb. An added custom force field is stored by GROMACS Wizard for later use.
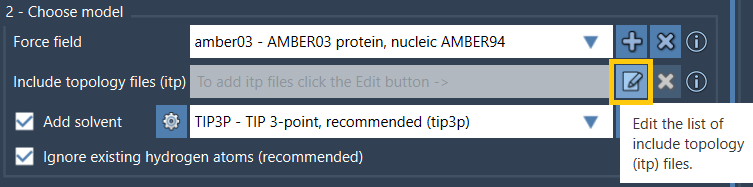
Before clicking Prepare, use this quick check: the selected system contains the intended ligand; the ligand name matches between structure and topology; the topology file belongs to the force-field choice; and any required custom force field has been added. Then complete the other preparation choices, such as solvent model, periodic box, and ions. If warnings appear, or you want to inspect what happened, open the GROMACS Wizard output.
When preparation succeeds, the resulting system is saved in the results folder. You can load it into SAMSON to inspect it, then continue with energy minimization, NVT and NPT equilibration, and production molecular dynamics.
For the full workflow and related guidance, see the SAMSON protein–ligand systems tutorial.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.