



A molecular system can look perfectly reasonable in a viewer and still be poorly prepared for a periodic simulation. The box is not just empty space around a protein: periodic boundary conditions mean that copies of the box repeat in every direction. If a molecule interacts with its own periodic image, the simulation can produce misleading forces. Choosing a box is therefore a practical modeling decision, not a cosmetic one.
In SAMSON’s GROMACS Wizard, you can fit a periodic box around the selected system, choose the unit-cell shape, and then set the amount of space around the solute. The useful question is not simply “How large should the box be?” It is “Which box-sizing approach fits this system and workflow, while leaving enough room for periodic boundaries?”
Start with the unit cell
The wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated-octahedron unit cells. For a straightforward preparation, choose a shape appropriate to your workflow, then use Compute fitted box to fit it to the selected system. The fitted box includes the atoms in the selection, with dimensions based on the system’s size in x, y, and z.
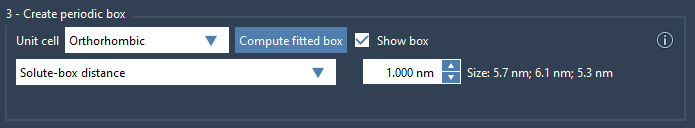
Before fitting, make sure the intended structures are selected in the Document view. If the preparation includes multiple structural models, the box must include all of them. A box fitted to only one structure may not contain the others.
Choose how the box is sized
After fitting, GROMACS Wizard offers two useful ways to define the box:
- Solute-box distance: specify the distance between the system and the box. The wizard fits the box around the solute using that spacing. The documentation recommends at least 1 nm to help satisfy the minimum image convention. For a batch project, each conformation or trajectory frame can have a different box size.
- Box lengths: specify the box dimensions directly. The initial fit is tight around the system, so you need to increase the dimensions to leave sufficient space. For a batch project, the initial box is determined from all conformations or the whole path, making this option useful when you want one box size throughout the batch.

These options answer slightly different needs. A solute-box distance is convenient when you want to define a consistent margin around each structure. Box lengths are useful when keeping the same dimensions across a batch matters more. Neither option removes the need to check that the resulting box is appropriate for the system and simulation.
Why the margin matters
Under periodic boundary conditions, the system is surrounded by repeated copies of itself. The minimum image convention is intended to prevent a molecule from interacting with its own periodic image within the simulation’s interaction range. The documentation recommends at least 1.0 nm between the system and the box edge. In practical terms, that corresponds to at least 2.0 nm between periodic images of a protein or other non-solvent molecule. Molecular structures can change conformation, so allow for more than the shape you see in the starting snapshot when needed.
With Box lengths, remember that a tight fit is only a starting point: increase the size to create the needed space. With Solute-box distance, set a suitable margin and review the generated dimensions. If you use a custom force field, check its publication or guidance for any specific cutoff-related requirements rather than assuming the general recommendation is sufficient in every case.
A quick preparation check
- Confirm the selection contains every structure you intend to prepare.
- Choose the unit-cell type and compute the fitted box.
- Pick solute-box distance for a margin-based fit, or box lengths when shared dimensions across a batch are important.
- Check that the box includes all selected structures and leaves adequate space for periodic images.
Once the box is defined, continue with the remaining preparation choices, such as solvent and ions, as required by your system. Note that adding ions requires adding solvent, because ions replace water molecules in the preparation workflow.
For the full preparation walkthrough, including force fields, solvent, ions, and running the preparation step, learn more in the GROMACS Wizard preparation documentation.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.