



A molecular dynamics run can fail before it begins—or proceed with a setup that does not match your intent. The preparation stage is where several consequential choices come together: which structure to simulate, how to describe its interactions, how much space to leave around it, and whether to add solvent or ions. Taking a moment to check these choices can save time later and make the resulting system easier to interpret.
This guide focuses on a practical question for molecular modelers: how do you prepare a system without overlooking the choices that can affect the simulation? The examples follow the GROMACS Wizard workflow in SAMSON, but the reasoning behind them is useful whenever you prepare a molecular system for simulation.
Start with the exact system you intend to simulate
Before choosing a force field or building a box, confirm which structural model is being prepared. In a document containing multiple models, select the intended structure in the Document view, then use Set system. If the selection is empty and multiple structural models are present, the wizard may ask whether to include them all.
This check matters especially when working with alternative conformations or a batch project. A batch can be useful when the same preparation and later simulation steps should be applied to a set of conformations, or when comparing related systems. Make the choice deliberately: a single system and a batch project are different workflows, even if they begin with similar structures.
If your system includes a ligand or another molecule that GROMACS cannot parameterize from the selected force field, have its topology files ready. Additional molecules may require .itp include files, and those files must be compatible with the force field used to parameterize them. A topology and force field that do not correspond can undermine an otherwise careful setup.
Choose the model as a compatible set
The force field is not just a dropdown choice: it defines how molecular interactions are represented, and its selection can influence the simulation. Consult the relevant force-field publications and choose one appropriate for your system. If using custom topology files, check that they were generated for the force field you intend to use.
For explicit-solvent simulations, choose a water model supported by the force field. In the tutorial example, the selected combination is OPLS-AA/L with SPC/E water. This is an example workflow, not a universal recommendation. If the water model is set to none, solvent and ions cannot be added through this preparation step.
Existing hydrogen atoms deserve a separate check. Their names must follow the conventions of the selected force field if you keep them. The wizard also provides an option to ignore existing hydrogens so GROMACS can add them using the selected force field’s naming convention. The documentation recommends this approach, particularly for NMR structures.
Give the system enough room
A periodic box is more than a container for solvent. Periodic boundary conditions mean the system is repeated in space, so an undersized box can bring a molecule too close to its periodic image. The minimum image convention is the reason to leave adequate space between the solute and the box boundaries.
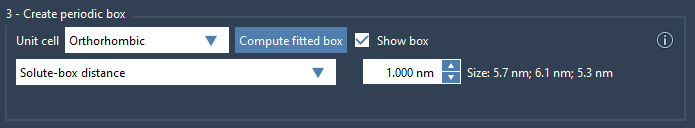
The wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated octahedron unit cells. You can fit a box using either Box lengths or Solute-box distance:
- Box lengths lets you specify the box dimensions and position. For a batch, the initial dimensions are determined from all conformations or the full path, which is useful when you want a consistent box size across the batch.
- Solute-box distance sets the distance between the solute and the box. The documentation recommends at least 1 nm to help satisfy the minimum image convention. For a batch, this can result in different box sizes for different conformations or trajectory frames.
For the tutorial, the suggested workflow is to choose an orthorhombic cell, compute the fitted box, and use the solute-box distance option. Whatever method you choose, ensure the box includes every structure selected for preparation. If you choose box lengths, remember that a tight fit is not automatically a sufficient simulation box: increase the dimensions as needed to leave space around the solute.

Add ions only after considering solvent
Neutralization is performed automatically by GROMACS when the Neutralize system option is selected. The total charge can be checked with the Update button. GROMACS adds positive or negative ions according to the system’s charge; it does not necessarily add both types.
Additional ions are a separate choice. You can specify a number of ions or a salt concentration. The documentation gives 0.15 M NaCl as a popular concentration representing physiological conditions, but the appropriate choice depends on the system and study. The concentration is calculated from the cell volume in the input .tpr file, and existing ions are not included in that specified concentration. Crucially, ions replace water molecules, so solvent must be added if you want to add ions.
Make the preparation easy to review
Choose a results folder where the generated project and later outputs will be saved. If useful, add custom index groups for later simulation or analysis; these can be generated from SAMSON selections, but the system needs unique, consecutive atom and residue indices. After clicking Prepare, check the output log and review the prepared structure when the import dialog appears. The resulting document includes index groups that can be selected for inspection.
A reliable preparation is a series of connected decisions: the correct structure, compatible parameter files, an intentional solvent and ion setup, and a box that accounts for periodic images. For the complete walkthrough, see the GROMACS Wizard preparation documentation.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.