



A molecular transition path can look plausible and still be a poor starting point for analysis: its intermediate structures may be unevenly spaced or may pass through geometries that are not energetically favorable. The Parallel Nudged Elastic Band (P-NEB) method in SAMSON helps refine a candidate path by optimizing its intermediate images while spring forces keep neighboring states distributed along the route.
The key is to start with the right expectation. P-NEB refines a path or a set of conformations you already have; it does not create the initial pathway between two states. If all you have are relaxed starting and ending structures, first generate intermediate conformations—for example, by linear interpolation or with a path-generation extension such as Ligand Path Finder. Then use P-NEB to improve that candidate route.
Choose a path when you can
SAMSON can apply P-NEB to either a path or a selected set of conformations. For the same system and number of steps, the documentation recommends using a path because optimizing a set of conformations takes longer. If your conformations are already in the document, you can combine them into a path by selecting them and choosing Conformation > Create path from conformations from the context menu.
A path stores a trajectory for a selected group of atoms. Conformations store saved positions for that group. This distinction matters in practice: a path is the convenient input when your candidate route is already represented as a trajectory, while a conformation set can be useful when your intermediate states are stored separately.
Prepare the model and settings
Before starting, add the P-NEB extension and the FIRE state updater, then load a path or conformations. The tutorial’s sample documents include a smaller Zinc ligand unbinding example and a lactose permease–thiodigalactoside complex with unbinding paths.
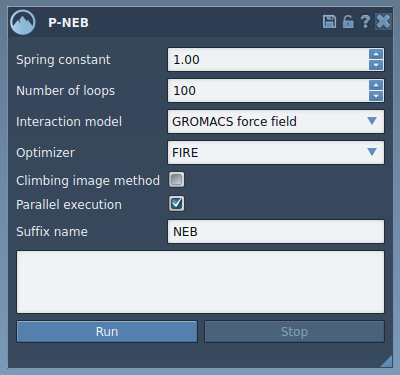
Open P-NEB through Home > Apps > All > P-NEB, or find it with the Find everything… search box. For the tutorial workflow, set the spring constant to 1.00, the number of loops to 100, choose Universal Force Field as the interaction model, and choose FIRE as the optimizer. Leave the climbing image option unchecked for the first run; you can repeat the calculation later with it enabled to try the climbing image strategy for locating a saddle point. Enable parallel execution, which runs one thread per conformation, and enter NEB as the suffix for the output name.
Run and inspect the result
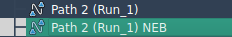
Select the candidate path in the Document view and click Run. When the Universal Force Field setup asks whether to use existing bonds, choose to use them and click OK. P-NEB will initialize and begin computing; progress is shown in the status bar. Once it finishes, a new path appears in the Document view, and the app displays a computation summary.

Do not treat completion as a substitute for inspection. Select the resulting path and examine it with the Inspector. You can also double-click it to start or stop its animation, making it easier to follow how the selected atoms move through the intermediate states. If you used separate conformations, the optimized conformations appear in the Document view and can be double-clicked to restore their corresponding positions.
In short: generate a candidate route first, prefer a path input when available, keep the force-field and optimizer choices consistent with your workflow, and inspect the optimized result before using it in downstream analysis. For the full walkthrough and related tutorials, learn more in the P-NEB documentation.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.