Selecting atoms with natural language in SAMSON AI
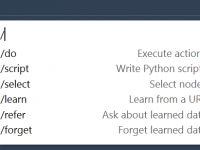
Molecular modeling involves frequent selection of specific atoms, residues, chains, ligands, or binding sites. This task can become tedious and error-prone, especially in complex molecular systems. Traditional selection tools involve dropdowns, checkboxes, or writing selection syntax. Not only does this…
Quickly Identify Bonds by Type in Your Molecular Models
Watching the SARS-CoV-2 Spike Move: A Useful Trajectory for Molecular Modelers
Why You Can Only Use One Editor at a Time in SAMSON
Avoid Solvent Clashes in GROMACS CG Simulations with One Simple Tweak
Filtering Side Chains by Atom Counts with NSL
Exporting DNA Nanostructures for Simulation with Adenita

A Simple Way to Record Molecular Motion Without Coding
Tracking how molecules move during simulations or interactive presentations can be key when communicating your discoveries. But if you’ve ever tried capturing atom trajectories while modifying structures in real-time—be it docking ligands, assembling molecules, or simulating dynamics—you might know how…