



Running GROMACS Simulations Without Interrupting Your Workflow
A Quick Guide to Covalent Docking in SAMSON with the FITTED Suite
Moving Groups of Atoms Without Losing Their Structure in SAMSON
How Embedded Python Scripts Make Molecular Modeling More Reproducible
Reducing False Positives in Protein Docking: Use Range Angles in Hex
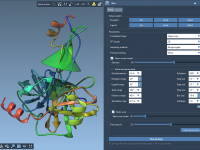
A Simple Way to Batch-Edit Molecular Attributes in SAMSON
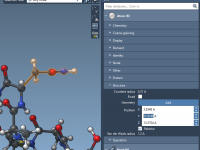
Quickly Filter Visible or Selected Property Models in SAMSON
Custom Index Groups in GROMACS Wizard: A Smart Way to Prepare for Targeted Simulations
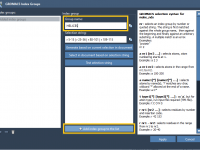
Cleaning Dozens of PDB Files Just Got Simpler: A Closer Look at Batch Protein Prepare
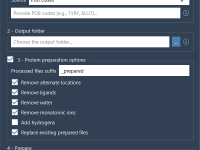
Anyone who has worked with multiple protein structures knows the challenge: each file needs cleanup before starting a molecular dynamics simulation, docking calculation, or structural analysis. Removing water, handling alternate locations, fixing missing hydrogens—it’s the sort of repetitive task that…