



A molecular system can look perfectly reasonable in a viewer and still be set up to interact with a copy of itself. That is one of the easy-to-miss consequences of periodic boundary conditions: the simulation treats the box as part of a repeating space, not as a container with an edge.
When preparing a solvated system, two questions help prevent a common setup problem: Which box shape fits the solute? And is there enough separation between the solute and its periodic images? The second question is especially important because an undersized box can make the forces on a molecule incorrect.
Think of the box as repeating
With periodic boundary conditions, the simulation represents neighboring copies of the unit cell. For short-range non-bonded interactions, GROMACS uses the minimum-image convention: it considers the nearest image of each particle. In practical terms, the solute should not interact with its own periodic image. Long-range electrostatics require additional methods, such as PME, but that does not remove the need to choose appropriate box dimensions.
A useful starting rule is to leave at least 1.0 nm between the solute and the box boundary. This gives at least 2.0 nm between periodic images of the solute. Treat this as a practical guide, not a substitute for checking the cutoff restrictions that apply to your simulation.
Choose a shape for the job
GROMACS Wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated octahedron unit cells. Cubes are familiar, but they are not automatically the most economical choice for every solute. For an approximately spherical macromolecule in solution, a rhombic dodecahedron or truncated octahedron can fit the shape more efficiently while maintaining a chosen distance to periodic images.
The rhombic dodecahedron is the smallest and most regular space-filling unit cell. At the same image distance, its volume is 71% of the volume of a cube. The documentation notes that this can save about 29% of CPU time when simulating a spherical or flexible molecule in solvent. The practical takeaway is not that one shape is always best, but that a shape suited to the solute can avoid filling unnecessary space with solvent.

Set dimensions deliberately
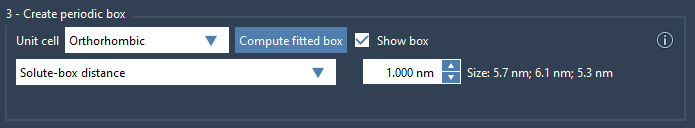
In GROMACS Wizard, the initial box can be fitted in two ways:
- Box lengths: Specify the box dimensions. The initial fit is tight, so increase the size as needed to provide enough solute-to-boundary distance. This option also lets you move the box to position the system. For a batch project, the initial size is based on all conformations or the whole path, which is useful when you want one box size throughout.
- Solute-box distance: Specify the distance between the solute and the box. At least 1 nm is recommended as a practical way to help meet the minimum-image requirement. In a batch project, box sizes can differ between conformations or frames.
A quick setup check
- Identify whether your solute is roughly spherical or has a shape that favors another box geometry.
- Choose a unit cell and decide whether consistent dimensions across a batch matter for your analysis.
- Check the solute-to-boundary distance; use at least 1 nm as a practical starting point.
- Confirm that the box dimensions are compatible with the simulation’s cutoff restrictions.
One detail can be confusing when inspecting results: GROMACS keeps particles in a brick-shaped volume for efficiency. When SAMSON loads GROMACS results, it attempts to detect the unit-cell type, and the type can also be changed in the trajectory importer dialog.
For the full explanation and links to the related preparation steps, see the GROMACS Wizard guide to periodic boundary conditions.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.