



A protein–ligand structure can look perfectly complete and still fail during simulation setup. A common source of trouble is that the ligand’s structure and its topology do not agree: atom names differ, required hydrogens are missing, or the topology depends on a force field that has not been supplied. These details are easy to overlook because they may not be obvious from viewing the complex.
This guide focuses on the handoff that often causes the most confusion: preparing a non-covalently bound ligand and connecting its parameters to the protein system in GROMACS Wizard. The key idea is to treat the ligand structure and its topology as a matched pair, then check that the force field used to parametrize the ligand is available when you prepare the complete system.
1. Start with the force field you intend to use
Before sending a ligand to a parametrization tool or server, decide which force field you plan to use for the simulation. The ligand parameters need to be compatible with that choice. The documentation lists examples: Antechamber for AMBER, ATB for GROMOS96 54A7, CGenFF for CHARMM, and LigParGen for OPLS-AA. These are examples, not interchangeable routes: choose the one appropriate to your planned force-field workflow.
Also check the input formats accepted by your chosen service before exporting the ligand. SAMSON can convert between file formats: load the structure and save it in a format accepted by the parametrization workflow.
2. Prepare the ligand before parametrization
Many parametrization tools expect the ligand to include its hydrogens. In SAMSON, hydrogens can be added through Edit > Add hydrogens. For ligands in the Chemical Component Dictionary (CCD), SAMSON can use the CCD’s hydrogen names. For ligands not in the CCD, hydrogen placement is based on valences, so aromatic rings and charges should already be specified. If a non-standard ligand file contains that information—for example, a .mol2 file—use it as the starting point where appropriate. External tools such as Open Babel are another option for adding hydrogens.
If the ligand is embedded in a protein–ligand structure, select it in the Document view and use Home > File > Save selection as… to export it. This gives you a separate ligand file to submit. If you already have a ligand file with hydrogens added, you can use that directly.
3. Keep the structure and topology in sync
After parametrization, collect the ligand’s include topology file, usually an .itp file. If the service provides an updated structure, use it to check whether atom names, hydrogens, or other structural details changed. This check matters: the ligand name in the structure must match the name used in the topology data you import.
If the parametrization service changed the ligand, do not assume the original ligand in the complex is still a suitable match. You can position the updated ligand at the old ligand’s location and delete the old one, or add the ligand by another suitable method. SAMSON supports manual placement with Move editors, docking with AutoDock Vina Extended, or appending the ligand structure to the protein structure file. When appending, ensure the structures do not overlap and use separate chains or consecutive sequence numbering.
4. Add the topology—and any required force field
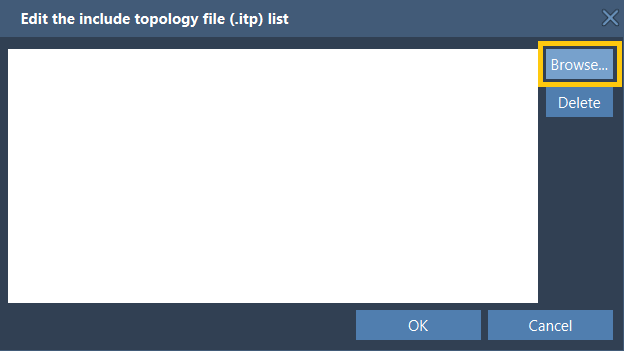
In GROMACS Wizard, open Home > Apps > Biology > GROMACS Wizard, go to Prepare, choose the protein–ligand system in Document view, and click Set system. In the Choose model section, use Edit beside the include topology files setting and browse to the ligand’s .itp file.
Some topology files rely on a custom force field that is not included by default. For example, an ATB-generated topology may require the corresponding gromos54a7_atb force field. Download the required force field from the service and add it using the force-field Add button. GROMACS Wizard stores an added force field for future use. The important practical check is to confirm both parts are present: the ligand topology and the force field it requires, if applicable.

5. Prepare, then continue the workflow
Complete the remaining preparation settings, such as the solvent model, periodic boundary box, and ions, following the general Prepare tutorial. Click Prepare to run the step. If a warning or other message appears, read it rather than dismissing it automatically; the documentation notes that warnings may occur, including when using the ATB GROMOS force field. You can inspect the run log with the Output button.
Once preparation succeeds, the resulting system is saved in the results folder, and you can optionally load it into SAMSON for inspection. From there, continue with energy minimization, NVT equilibration, NPT equilibration, and production molecular dynamics.
This workflow covers non-covalently linked complexes. Covalent protein–ligand links need additional treatment for parametrizing the linking bond, so they are outside the scope of these steps. For the full instructions and related details, learn more in the original protein–ligand systems tutorial.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.