



Choosing a simulation box can feel like a geometry problem with consequences you only discover later. Make it too small, and the solute may interact with its own periodic image. Make it unnecessarily large, and you add solvent molecules that must be simulated. The goal is to choose a box shape and size that suit the system while respecting the minimum image convention.
Why the box matters
GROMACS applies periodic boundary conditions: the simulation cell is repeated in every direction. For short-range non-bonded interactions, the minimum image convention means that the nearest image of each particle is considered. If a solute is too close to the edge of its box, a periodic copy can become close enough to affect it. The resulting forces may then be wrong.
Long-range electrostatics need additional care. GROMACS can use lattice-sum methods such as Ewald summation, PME, and PPPM, but this does not remove the need to choose box dimensions that meet the relevant cutoff restrictions.
Start with the shape
GROMACS Wizard supports cubic, orthorhombic, triclinic, rhombic dodecahedron, and truncated octahedron unit cells. The best choice depends on the system and how you want to fit it.
For an approximately spherical macromolecule in solution, the rhombic dodecahedron and truncated octahedron can be useful alternatives to a cube. They are more sphere-like, so they can require fewer solvent molecules for a given minimum distance between the macromolecule and its periodic images. The rhombic dodecahedron is the smallest regular space-filling unit cell; at the same image distance, its volume is 71% of that of a cube.

That volume comparison is about geometry, not a guarantee of a particular runtime for every model. The practical benefit is that a more suitable cell shape may reduce the amount of solvent needed while preserving the desired distance to periodic images.
Then set the distance
A practical rule of thumb is to leave at least 1.0 nm between the solute and the box boundary. This gives at least 2.0 nm between periodic images of the solute. Treat this as a starting point, not a replacement for checking the cutoff restrictions relevant to your simulation.
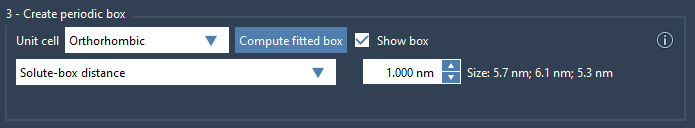
In GROMACS Wizard, you can approach the fit in two ways:
- Box lengths: specify the dimensions of the box. The initial fit is tight, so increase the size as needed to provide sufficient separation. This option also lets you move the box to position the system. For a batch project, the initial size is based on all conformations or the entire path, which is useful when you want one box size throughout.
- Solute-box distance: specify the distance from the solute to the box. At least
1 nmis recommended as a practical starting point for the minimum image convention. In a batch project, box sizes can differ for each conformation or path frame.
A quick decision checklist
- Consider the solute’s shape. For an approximately spherical molecule in solvent, compare the rhombic dodecahedron or truncated octahedron with a cubic box.
- Choose whether a fixed box size or a solute-to-box distance better suits your setup, especially if you are preparing multiple conformations.
- Check the separation between the solute and its periodic images. Use the
1.0 nmsolute-to-boundary guideline as a practical starting point, and consult the GROMACS cutoff restrictions for your simulation.
One detail that can be surprising: GROMACS keeps particles in a brick-shaped volume internally for efficiency. When you load GROMACS results, SAMSON attempts to detect the unit-cell type, and the importer dialog lets you adjust it if needed.
For the full explanation and setup context, see the GROMACS Wizard guide to periodic boundary conditions and Step 1: Prepare.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.