Author: OneAngstrom
Quickly Filter Visible and Hidden Elements in Molecular Models
How to Keep Functional Waters and Delete the Rest in Biomolecular Simulations
Prevent unwanted camera drift when animating molecular systems
When creating animations of molecular systems, you’re often focused on illustrating detailed transitions—like conformational changes, ligand docking, or energy minimizations. But there’s a subtle issue that many researchers encounter: the camera viewpoint can drift between frames unintentionally, especially if no…
Streamline Your Molecular Workflow with SAMSON AI’s Python Scripting
Writing Python scripts for molecular modeling often involves switching between documentation, coding environments, and visualization tools. For researchers and students working with complex simulations, simple tasks—like transforming atoms or building interfaces—can become time-consuming due to repetitive coding and context switching.…
Quickly Select Atoms by Geometry with NSL in SAMSON
When modeling molecules, filtering atoms by chemical properties or structural features is an everyday task. Yet, for many molecular modelers, selecting atoms by geometry—like tetrahedral or octahedral configuration—can still be a tedious, time-consuming process, especially when working with large systems.…
Exporting Ligand Trajectories for Free Energy Calculations in SAMSON
Refining Ligand Unbinding Pathways: A Step-by-Step Guide Using P-NEB in SAMSON
Before You Simulate: The Smart Way to Clean Up Molecular Geometries
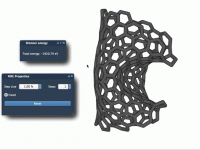
When preparing molecular systems for simulation, clean geometry isn’t optional—it’s essential. Poor atomic arrangements can lead to unrealistic behavior, convergence failures, or wasted compute time. That’s why optimizing geometries before launching simulations should be part of every molecular modeler’s workflow.…