



Two protein structures can show different conformations, but comparing only the endpoints leaves a practical question unanswered: what might the structural change look like in between? Building a sequence of intermediate conformations by hand is tedious, and simply comparing the two structures does not provide a continuous path to inspect or export.
SAMSON’s ARAP Interpolator can generate a smooth geometric transition between two protein conformations. This can help you inspect a structural change, prepare a path for downstream workflows, or create intermediate structures for further analysis. The key is to prepare and match the inputs carefully: interpolation is only useful when the structures represent the intended corresponding parts of the protein.
Start with comparable structures
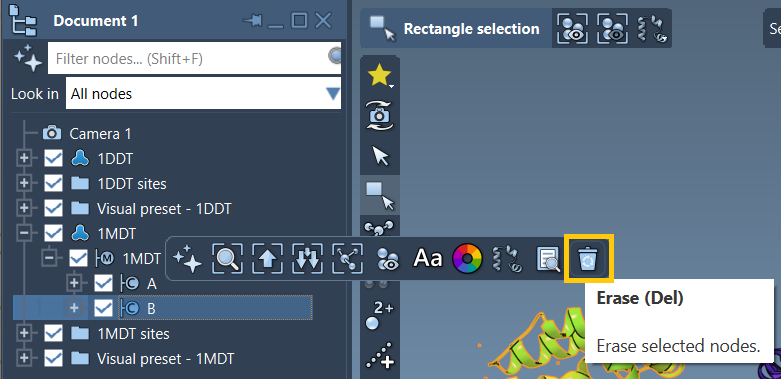
The tutorial uses two conformations of diphtheria toxin: PDB structures 1DDT and 1MDT. In SAMSON, fetch both structures, then work with chain A in each. The example removes chain B from 1MDT, cleans both structures with Home > Prepare, and creates a named conformation for each structure.
This preparation is more than housekeeping. Extra chains, water, ions, or ligands can leave a structure with disconnected components, which may prevent the interpolation from running. It also helps to be deliberate about which chains you are comparing: the start and goal should represent the parts of the system you actually want to connect.
Choose how atoms and connections are represented
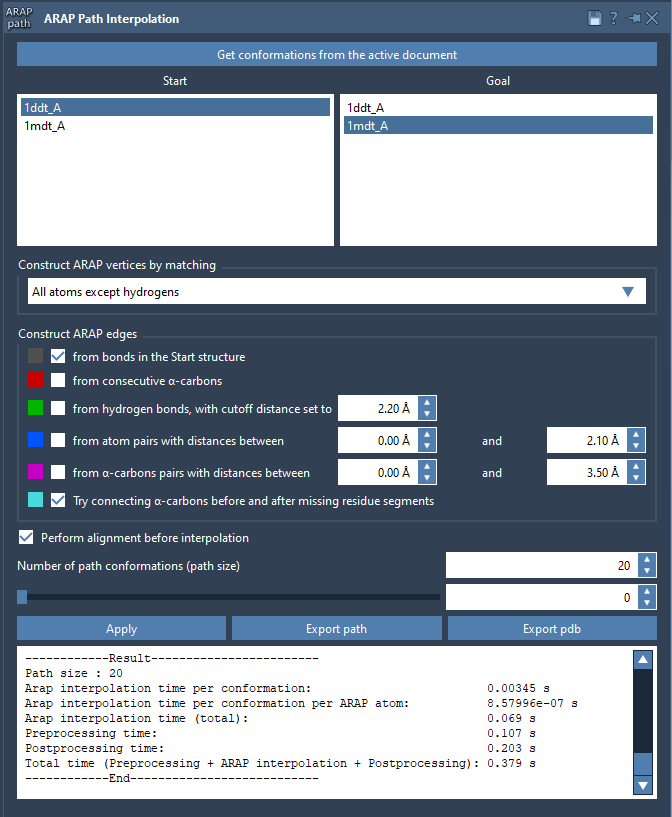
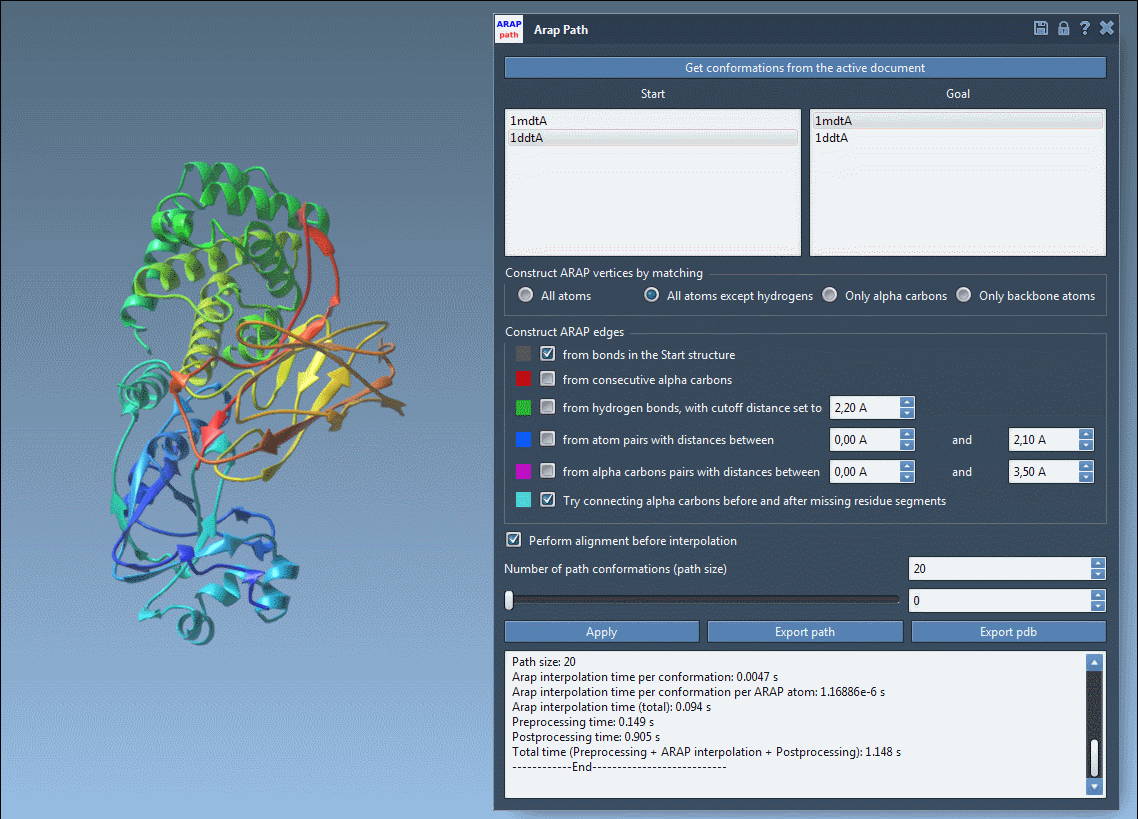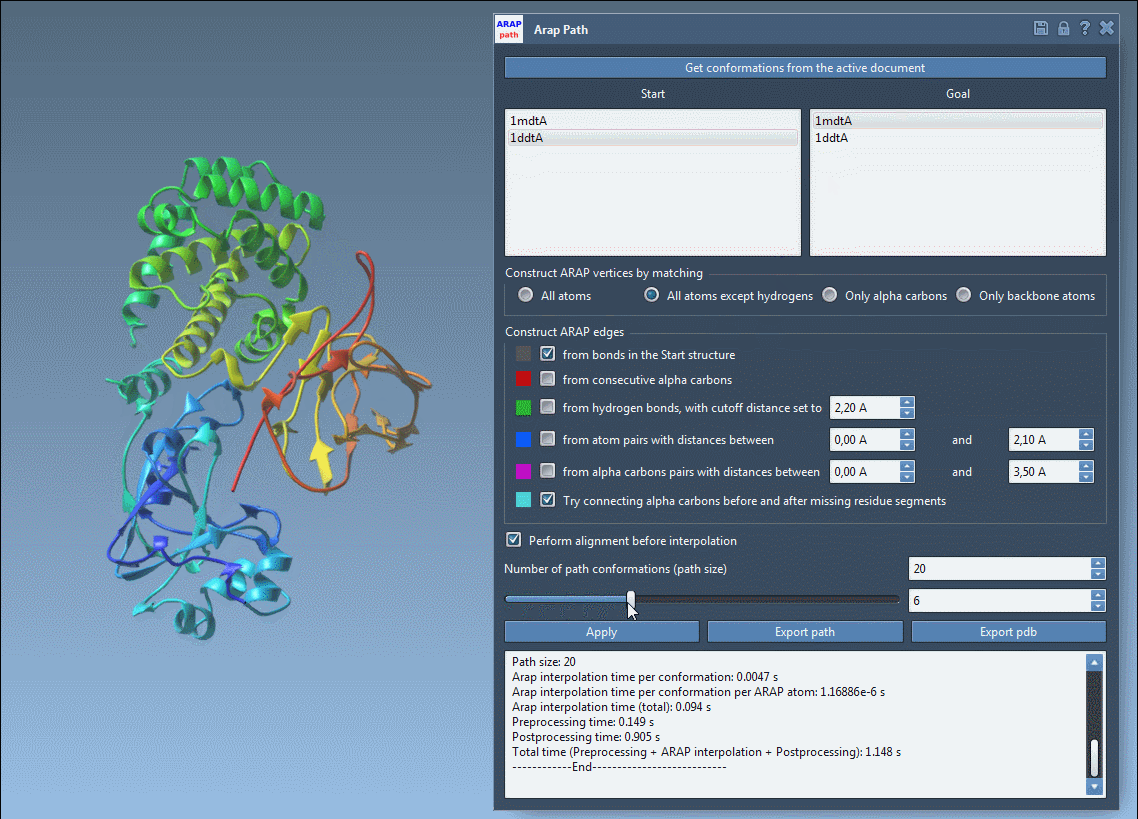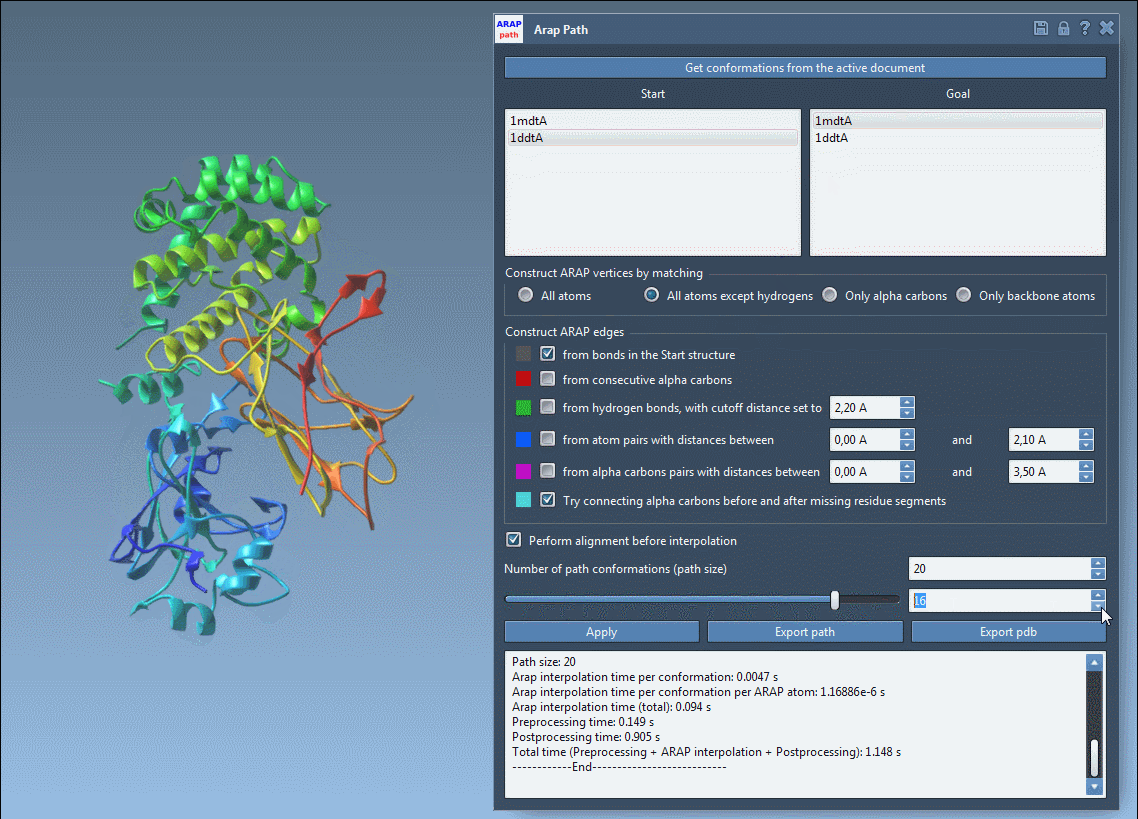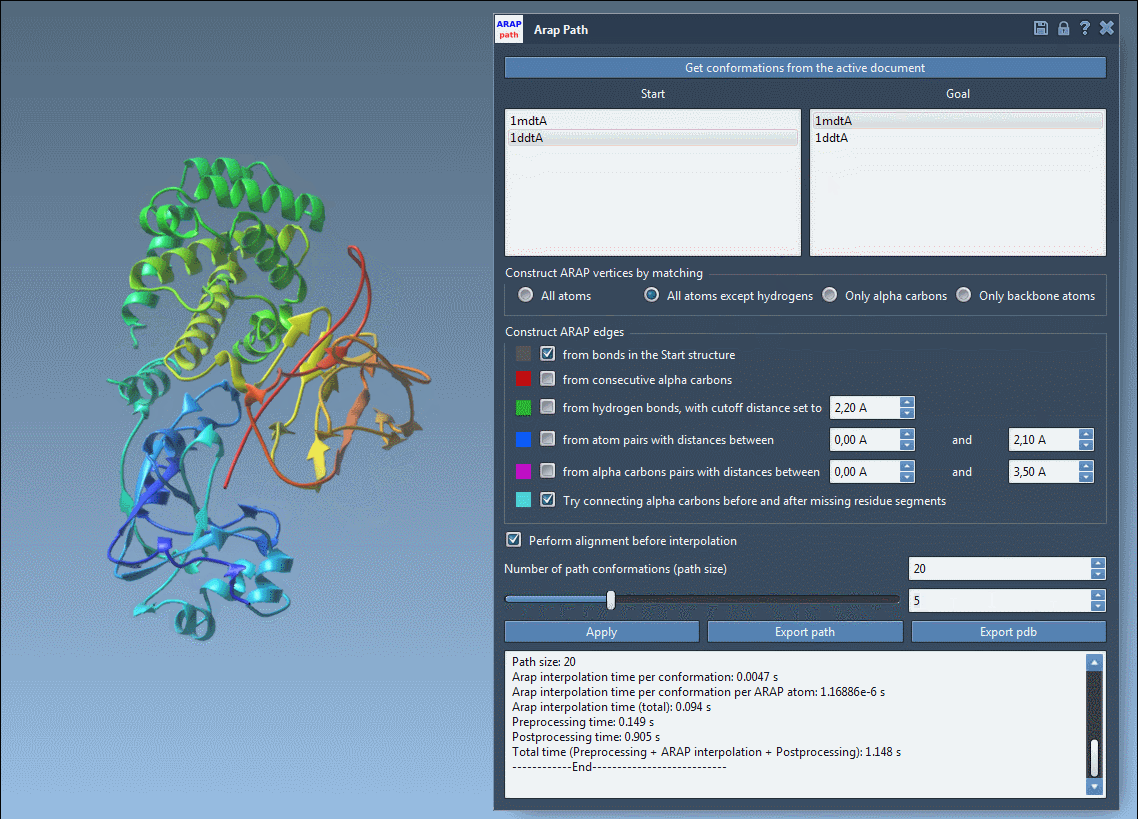
In the ARAP Path Interpolation app, load conformations from the active document and select the start and goal. The example uses 1DDT A as the start and 1MDT A as the goal.
For atom matching, the tutorial recommends All except hydrogens. This matches non-hydrogen atoms between the conformations and uses the matched start atoms as ARAP vertices. Other available choices include all atoms, only α-carbons, or only backbone atoms. The appropriate choice depends on what level of structural detail you need to represent.
Connectivity also affects the result. For the example, enable from bonds in the Start structure and Try connecting α-carbons before and after missing residue segments. The first uses covalent bonds in the start structure to create ARAP edges; the second adds connections across missing-residue segments. The app also offers other edge-construction methods, such as consecutive α-carbons or distance-based pairs. When multiple methods are selected, edges are added in the order shown in the interface.
Align first, then inspect the path
Enable Perform alignment before interpolation so the start conformation is aligned with the goal before the path is computed. Set Number of path conformations to 20 to include 20 conformations total, including the start and goal. Then run the interpolation.
After computation, use the slider to move through the conformations rather than judging the result from its endpoints alone. Scrubbing the path can help reveal where a motion is concentrated and whether the intermediate structures are visually coherent. The app also creates an ARAP edges visual model; show or hide it in the Document view to inspect how the chosen connectivity contributes to the construction.
Know what the path does—and does not—tell you
An interpolated path is a useful geometric representation of a transition, but it should not automatically be treated as a validated physical mechanism or a free-energy result. Inspect the intermediates and use the path as an input for an appropriate next step. The tutorial points to umbrella sampling with the GROMACS Wizard, steered molecular dynamics, or refinement with P-NEB as possible follow-on workflows. You can export the path as a trajectory in the current SAMSON document or export the path and its conformations to PDB files.
If the computation reports that the structure does not make one connected component, check both inputs for unremoved water, ions, ligands, or other disconnected structures, then run Home > Prepare again.
For the full walkthrough and additional options, see the ARAP protein interpolation tutorial.
Note: SAMSON and all SAMSON Extensions are free for non-commercial use. Get SAMSON.