



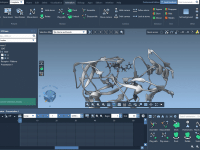
Streamline Your Molecular Structures with the FIRE Minimizer
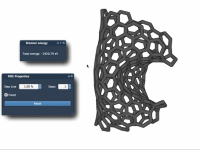
In molecular modeling, achieving stable and realistic molecular structures is crucial for simulations and workflows. One of the most common challenges modelers face is quickly optimizing complex geometries, especially when dealing with large-scale structural motions. The Fast Inertial Relaxation Engine…
Simplify Molecular Dynamics with GROMACS Wizard’s Custom Parameters
Efficient Filtering with “structuralModel” Attributes in SAMSON
Streamlining NVT Equilibration with GROMACS Wizard: A Beginner-Friendly Guide.
Make Your Molecular Models Self-Contained with File Embedding
Easily Navigate File Attributes in SAMSON’s Node Specification Language
Tracking Molecular Interactions with the Contacts Analysis Tool in SAMSON
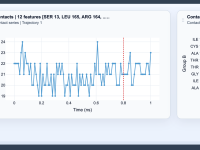
Easily Track Molecular Motion with SAMSON’s Distance Analysis
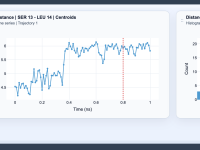
For molecular modelers, the ability to measure and analyze distances between selected groups in a molecular system is invaluable. Whether you’re investigating domain motions, tracking binding site transitions, or monitoring specific contacts during a molecular trajectory, understanding these distance-based dynamics…
Understanding and Utilizing Average Velocity in Molecular Dynamics.
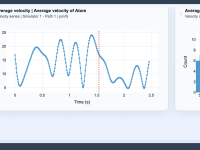
Molecular modelers and computational scientists commonly grapple with the challenge of analyzing dynamic trajectories effectively. Whether you are identifying areas of rapid movements in a molecular system or tracking moments of relative calm, understanding velocity changes across a time-based trajectory…