Streamline Your Molecular Design: Geometry Relaxation While Editing
For molecular modelers, creating and refining molecular structures often involves precise adjustments to geometries. A common challenge is ensuring that these modifications don’t compromise the stability or realism of your system. SAMSON simplifies this process by offering integrated geometry relaxation…
Simplify Molecular Presentations with the Disappear Animation
Building a Public Profile on SAMSON Connect.
Streamlining Molecular Modeling with NSL Selection Expressions
Molecular modeling often involves selecting specific atoms, residues, or other molecular components for analysis or manipulation. Whether you’re identifying atoms within a binding pocket or pinpointing residues involved in key interactions, accurate and efficient selection methods are critical. This is…
Unveiling Molecular Stability with Contact Persistence
How to Effortlessly Perform GROMACS Simulations in the Cloud
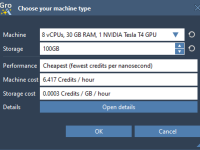
Molecular modelers often encounter a common challenge: limited computational resources on personal machines, especially when tackling large and complex molecular systems. Running GROMACS stages like equilibration or production molecular dynamics locally can be a significant bottleneck. This is where the…
Effortlessly Track Molecular Movements with the ‘Follow Atoms’ Animation
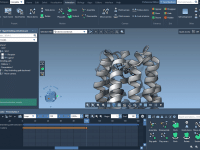
Simplifying Molecular Visualizations with Light Attributes in SAMSON
For molecular modelers and researchers, having clear and flexible control over visualization parameters can make a big difference in your workflow. Whether you’re crafting publication-ready figures or navigating complex molecular assemblies, the ability to fine-tune light attributes offers a significant…